Convergence: EU officials tout pending pharmaceutical reforms
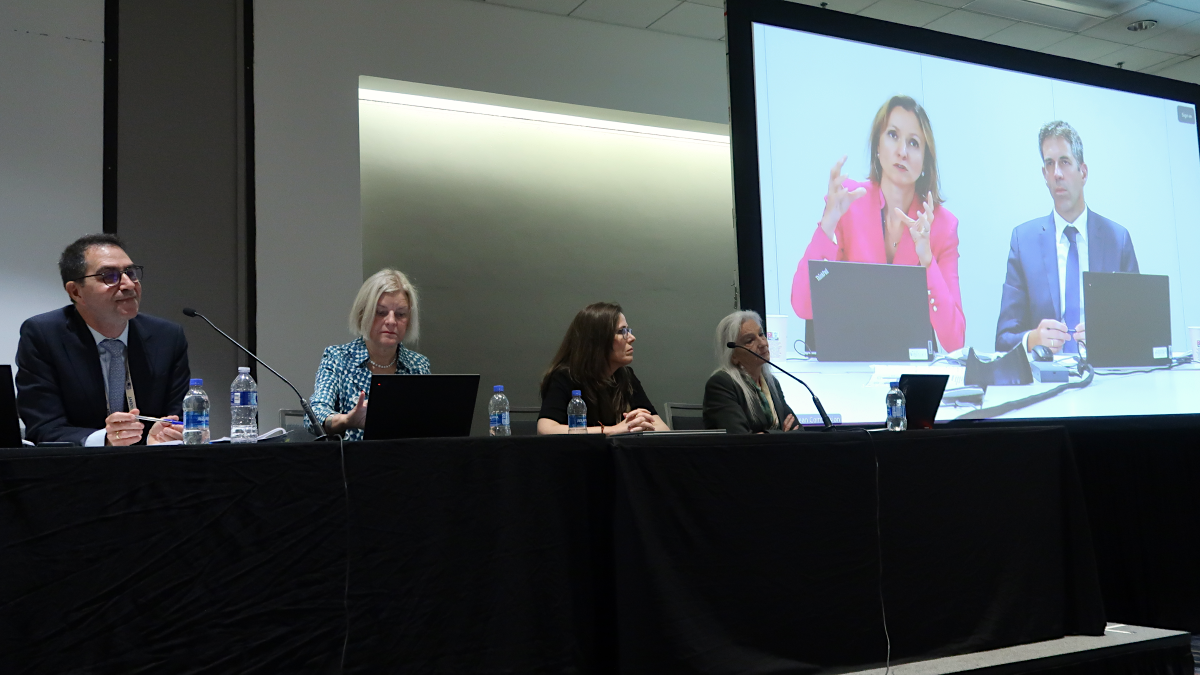
Left to right: Alberto Gañan Jimenez, Sabine Haubenreisser, Claudia Dollins, Andrea Laslop, Lilia Luchianov, and Antonios Rodiadis. (Credit: Ferdous Al-Faruque)
CHARLOTTE, NC — Officials from the European Commission and European Medicines Agency (EMA) touted key reforms presented in the proposal to overhaul the bloc’s pharmaceutical legislative framework during a panel discussion at RAPS Convergence 2026.
Antonios Rodiadis, a policy officer at the European Commission's Directorate-General for Health and Food Safety (DG SANTE), noted the reforms proposed in the new EU pharmaceutical legislation are intended to revise two decades of EU pharmaceutical rules. It includes a new Directive and Regulation to update and simplify the region's pharmaceutical rules and regulatory framework and added that the reforms are also intended to complement the proposed Critical Medicines Act, Biotech Act, Medical Device Regulation (MDR), and more.
"It's not just a reform, actually, you could call it a structural shift of EU pharma policy," said Rodiadis. "Because really, what it does is address the challenges that we've been seeing in the EU context in the past years, whether it relates to access to medicines.
"Even within the European Union, there's been a huge discrepancy in certain cases between member states' access to even centrally authorized medicines," he added.
Rodiadis noted that in 2025, certain larger member states had access to 80% of authorized drugs, while smaller member states only had access to less than 20% of them. He also noted that under the current EU regulatory system, it takes 430 days to authorize a new drug, which was exacerbated during the COVID-19 pandemic.
Rodiadis said those are some of the issues the new pharmaceutical legislation aims to address, among others. Some of the reforms he highlighted include reforming the EMA committees from a six-committee structure to a two-committee structure.
"This radically simplifies the system because you don't need more," said Rodiadis. "You need one committee for human medicinal products for the authorization process for everything that comes and is new, and then another, second committee, for the pharmacovigilance activities, the [Pharmacovigilance Risk Assessment Committee (PRAC)].
"This achieves simplification in the sense that, for example, today if you would have a product that is a pediatric rare medicine product using some kind of advanced therapy mechanism, it would have to go actually through five committees," he added.
The proposed reforms also include streamlining regulatory procedures, such as eliminating authorization renewals that created lags in the regulatory system; conducting phased reviews of data for drugs of major public health interest, which comes from EMA's experience conducting rolling reviews during the COVID-19 pandemic; and reducing the time for scientific assessments from 210 days to 180 days. For accelerated scientific assessments, Rodiadis said some drugs could be reviewed in 150 days.
"The simplification aspects of the system we have calculated will result into a €300 million annual reduction of costs, both for the authorities, public authorities, but also for industry," he added.
Rodiadis said the proposed legislation will also make pharmaceutical regulation more digital, and data driven. He noted that regulatory agencies and national regulators would be able to tap into health registries such as hospital databases and biobanks for regulatory purposes that may ultimately be used for marketing authorization and pharmacovigilance. He also said it would allow electronic submission of premarket applications and let manufacturers provide product information in only electronic format.
Additionally, Rodiadis said the proposed reforms would allow regulatory sandboxes so regulators can test regulatory innovations when considering bringing groundbreaking therapies to market and would provide vouchers to prioritize new antimicrobial drugs.
"This is actually an unprecedented regulatory tool," said Rodiadis. "Our analysis has shown that it can actually provide a very serious incentive to essentially provide funding and access to venture capital to those antimicrobial developers.
"We're talking about developers of not just any new antimicrobial, but really antimicrobials that are addressing multi-drug-resistant organisms," he added. "It's a new class or a new mechanism of action, for example, and what the voucher does is essentially it provides an additional year of data protection to the antimicrobial developer that can then be sold on the private market."
Rodiadis said the reforms would allow the use of adapted frameworks for novel medicines, provide early regulatory support through the European Medicines Agency (EMA), allow authorization of platform technologies such as messenger RNA vaccines, allow for decentralized manufacturing so that patients can access products closer to home, and provide more clarity for products that come under multiple regulatory frameworks.
Lilia Luchianov, who is also a policy officer at the European Commission's DG SANTE, provided insight into how the reforms would go into force if approved and implemented by the European Parliament and European Council. She said reforms such as the antimicrobial voucher, regulatory sandboxes, EMA international regulatory cooperation, and awarding certain EMA grants would go into effect immediately. However, monitoring and shortage management provisions, access provisions for new medicines, and other provisions would be phased in between six months and two years after the law is enacted to give manufacturers more time to transition.
Luchianov noted that they've made a concerted effort to ensure the new pharmaceutical legislation is logically aligned with other pieces of legislation, including the Biotech Act, in specific areas such as clinical trials and borderline and combination products. She said the new pharmaceutical legislation also tries to align with the topic of regulatory sandboxes.
"Our legislation foresees that when there is an authorization for clinical trials that is covered by a pharma sandbox, then there has to be a coordination," said Luchianov. "This has now been organized so everything happens smoothly, including the exchange of information between the authorities dealing with the sandbox, but also including all the necessary learning because the sandbox is definitely a learning exercise for the authorities and the companies involved in the development of that given product."
EMA preparation
Sabine Haubenreisser, EMA principal scientific administrator for stakeholders and communication, said EMA and the European medicines regulatory network are already preparing to implement the proposed legislation. She echoed that the reforms proposed in the legislation would lead to a simpler regulatory environment, provide more regulatory support, improve conditions for innovation, strengthen safeguards against drug shortages, and incentivize the development of antimicrobial drugs.
Haubenreisser noted that the proposed reforms would not only simplify the expert committee structure but also strengthen it. She said the experts from the disbanded committees would still be available to support committee work, and the committees would have formal representation from patients and healthcare professionals who would be voting members.
"In order to really be successful, we have been reviewing the text from day one," said Haubenreisser. "We have done a gap analysis and really identified what are really the actions and deliverables.
"We performed a resource allocation for the agency, but also for the network, and in sync with the governance structure ... we have identified the priorities for the next couple of months," she added.
Haubenreisser also said regulators will convene a multi-stakeholder workshop as soon as the new pharmaceutical legislation is approved and published to review the reforms and gather feedback.
Industry perspective
Claudia Dollins, GSK VP for precision medicine and global regulatory affairs, spoke on behalf of the European Federation of Pharmaceutical Industries and Associations (EFPIA) and emphasized that the proposed reforms were the largest change in the EU regulatory framework in two decades.
"This is a profound change, and it's a profound change in how medicines and vaccines will reach patients going forward," said Dollins. "It's an exciting time in the industry."
Dollins said the pharmaceutical industry has questions and concerns about the new pharmaceutical legislation, including concerns that guidances and assessment requirements may be issued without sufficient advance notice. She said there needs to be reasonable, feasible timelines for industry to implement the changes, and that they also need to be pragmatic. She also said there needs to be clarity regarding which products fall within the scope of different provisions and how to plan to meet those provisions.
"Really, we are looking for seamless integration and seamless transition into the physical framework," said Dollins. "What you have seen is a number of steps already being considered, [such as the] phased approach to implementation, to ensure that this is going to be the case.
"This is an exciting time with more than 100 implementing actions ahead," she added. "Industry stands ready to contribute expertise, data, and pilot experience at every step."