FDA officials detail inspection expectations for combination products
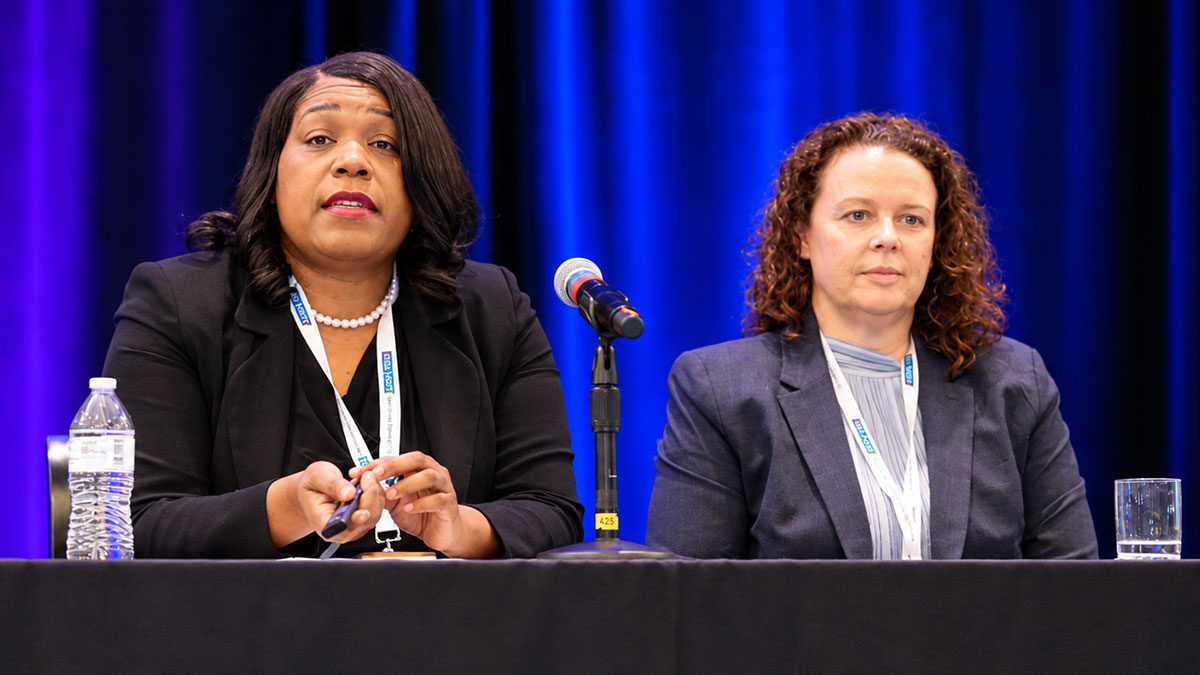
BALTIMORE – US Food and Drug Administration (FDA) officials said that manufacturers who opt to follow a streamlined approach to their combination product current good manufacturing practice (CGMP) should be aware that the agency will closely scrutinize their risk management activities at the AFDO/RAPS Healthcare Products Collaborative Combination Products Summit on Wednesday.
FDA’s streamlined approach to combination product CGMP compliance allows manufacturers of single-entity and co-packaged combination products to use a single quality system based on either drug CGMPs or the Quality Management System Regulations (QMSR), with specified provisions from the other framework incorporated to ensure compliance with requirements with the combination product CGMP requirements under 21 CFR Part 4.
Keisha Thomas, associate director for compliance and quality at the Center for Devices and Radiological Health (CDRH), emphasized that manufacturers of combination products should be aware of the new device “callout” provisions under QMSR and what to expect during inspections.
The QMSR made conforming edits to 21 CFR Part 4, to note specific quality provisions that a manufacturer must meet when using the streamlined CGMP pathway outlined in 21 CFR 4.4(b)(1). Instead of referencing the outdated Quality System Regulation (QSR) sections, this provision now aligns those callouts with the clauses of ISO 13485:2016.
Examples of products that qualify for these callouts include prefilled syringes with a drug, drug-filled autoinjectors, drug-containing inhalers, and medical devices that are drug-coated or drug-eluting.
As a result of the changes under QMSR, one of the things that FDA will be paying more attention to is the top management’s involvement in quality management. “The take-home message here is that one of the fundamental shifts with QMSR has been the extra emphasis on management responsibility in ensuring that quality management system requirements are met. It is not enough to ensure there is a quality management system. It is not enough to have quality documentation. It is imperative that top management play an integral role in this.”
Karen Masley-Joseph, senior advisor in the Office of Medical Devices and Radiological Health Inspection (OMDRHI) at FDA, added that investigators will be focusing on whether companies have adopted risk management activities throughout the product lifecycle.
“During an inspection, each and every time investigators come out, they will review your risk management documents.” She added that the risk management documents are “living documents” that should continue to evolve throughout the product’s lifecycle.
“The risk management document should not be the same one you submitted to the agency years before; it is a living document, and we will be requesting it each and every time. We are going to look at that risk management to see what hazards and harms you have mitigated,” she said.
Investigators will also be looking at purchasing controls and whether manufacturers are adequately controlling their supply chains. “Supply chain is a really big part of where risks can happen,” Masley-Joseph added.
She mentioned that there is an expectation in 21 CFR 4.4(b)(1) under clause 7.4 that specifies the criteria for evaluating your supplier. “A one-size-fits-all approach will not cut it for suppliers. Different suppliers have different risks in what they are supplying.”
Another change is that during inspections, the FDA can now review internal audit reports, supplier audit reports, and management review records. Under the former Quality System Regulation, investigators were not allowed to request these reports. “That exception is gone in the QMSR. It no longer exists,” Masley-Joseph said.