Why FDA finds Form 483 responses inadequate
Most companies treat the Form 483 response as the chance to close the loop after an FDA inspection. In many warning letters, FDA reaches a different conclusion: the company responded, FDA reviewed the response, and the agency still found the answer incomplete.
Atlas Compliance reviewed 267 FDA warning letters issued between January 2025 and August 2026. In 224 of those letters, or 83.9%, FDA found at least part of the company’s earlier response inadequate. Across the set, Atlas Compliance tagged 671 rejected-response findings and grouped them into 193 compliance themes.
Based on that analysis, FDA rarely names the problem for the first time in a warning letter. In many cases, the company answered the inspection finding but left the agency’s next question unresolved.
FDA looks for evidence behind the corrective action
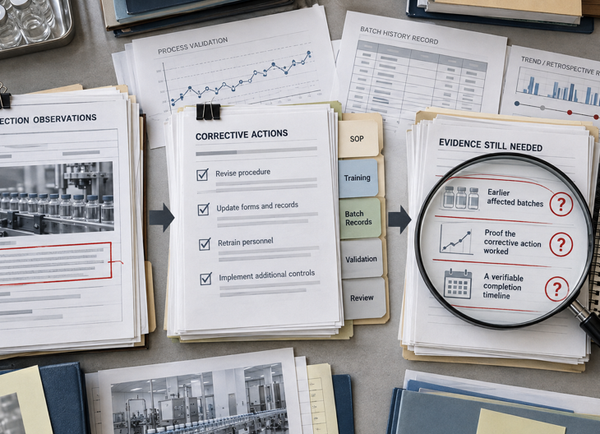
Many companies recalled product, opened corrective and preventive actions (CAPAs), revised procedures, submitted records, or wrote to FDA more than once. FDA still found responses inadequate when those actions did not demonstrate that the company understood the full risk, addressed earlier work, or verified that the correction worked.
In a 2025 warning letter involving a compounded pharmaceutical company, FDA credited corrective actions after the firm recalled injectable lots and responded to inspection findings. FDA still found the response inadequate because the firm had not evaluated the contamination risk from a textured wall panel near sterile production or explained how it would control that risk.
Another letter involving a ventilation device manufacturer in Hong Kong demonstrates the same issue within a different quality system. FDA investigators found that customer complaints were arriving by email without formal logging or review. The company submitted a CAPA report and revised its complaint-handling procedure. FDA still found the response inadequate because the company did not explain how it would handle complaints processed under the old system and did not show that staff had been trained on the revised procedure.
In both cases, FDA evaluated the risk left by the previous process. The response needed to account for work already completed under that process and show how the company would control the risk going forward.
Inadequate responses appear across product types
The pattern was strongest in drug warning letters. In the review, 87.8% of human drug warning letters included at least one inadequate response. The rate was 85.7% for biologics letters and 69.6% for medical device and radiological health warning letters.
FDA’s evidence requests varied by product type. Drug and biologics responses often stand or fall on batch records, laboratory data, monitoring results, contamination controls, and validation evidence. Device responses more often turn on whether the company can show that its quality-system process worked as written.
Across the full data set, the most frequently rejected response themes were document control, contamination control, process validation, data integrity, record review or retrospective assessment, deviation investigation, specification control, stability program, training and qualification, and aseptic processing.
Document control and contamination control accounted for 79 tagged response failures.
Procedures are a common weak point
Atlas Compliance also reviewed recurring language in the findings on rejected responses. One word appeared more often than any other: procedure.
"Procedure" appeared in 38% of rejected-response findings. "Oversight" appeared in 21%. "Validation" appeared in 19%. "Documentation" appeared in 12%. "Investigation" appeared in 10%. "Training" appeared in 7%.
These words describe companies whose quality systems existed on paper but did not hold up under FDA review.
A revised standard operating procedure (SOP) addresses the document. FDA may also ask whether the old procedure failed elsewhere or if the affected product remains in distribution. The response may need to show quality-unit oversight, evidence for the root cause, and proof that the fix worked.
The missed question is often retrospective
One recurring issue in the data was the look-back. A company may answer the specific observation in front of it, but miss the broader population that FDA is concerned about.
If FDA cites one batch, the response may need to address similar batches. If FDA cites one complaint file, the response may need to address complaints handled under the same process. If FDA cites one unvalidated step, the response may need to address product released before the validation work is complete.
A response can appear complete internally and still fail under FDA review when it answers the finding without addressing the risk around it.
Several warning letters in the review showed this problem at scale. One company had responses flagged across nine unrelated themes in a single letter, including Design History File remediation, equipment qualification, facility maintenance, and an out-of-specification (OOS) investigation. Another was also flagged across nine themes, including complaint handling, design control, UDI compliance, and a post hoc validation study.
One inadequate answer may reflect a documentation gap. When FDA finds gaps across nine unrelated themes in one letter, the pattern can indicate that the CAPA package moved faster than the investigation behind it.
Five questions before sending a response
Before submitting an inspection response, quality and regulatory teams can test it against five questions drawn from the gaps in the Atlas Compliance review.
- What happens to product already made, shipped, tested, or released while the fix is still underway?
- How far back did the company look? A single observation may point to a wider population of batches, complaints, tests, records, suppliers, or devices.
- What evidence supports the root cause? A conclusion without supporting data, investigation records, or ruled-out alternatives may read like no root cause at all to FDA.
- What proves the fix worked? Monitoring results, requalification data, repeat testing, or effectiveness checks may matter more than the promise to revise a procedure.
- What dates can FDA verify? An open-ended commitment gives the agency little to evaluate.
These questions map closely to gaps that FDA identified repeatedly, although no checklist can guarantee that FDA will accept a response.
Patterns emerge across warning letters
One warning letter shows what went wrong at one company; however, by searching a tagged collection, quality teams can see which mistakes repeat across product types, sites, and response themes.
Before FDA reviews its own response, teams can search comparable letters by product type, keyword, site, company, investigator, Code of Federal Regulations (CFR) section, or response theme.
Atlas Compliance structures FDA inspection and enforcement data so teams can search across warning letters, Form 483s, Establishment Inspection Reports (EIRs), observations, investigator profiles, and global enforcement records from regulatory agencies.
For this analysis, Atlas Compliance linked each rejected response to the company, product category, response language, keyword, what FDA asked for, and the source warning letter. That structure turns public enforcement records into a searchable tool for preparation.
In many warning letters, FDA identified a second failure: the company had answered the inspection finding, but its response did not resolve the agency’s concern.
See how Atlas makes the FDA enforcement record searchable across warning letters, Form 483s, EIRs, observations, and response themes before your team drafts its next response.
Bhavish Agarwal is the founder and CEO of Atlas Compliance; he previously worked at Google and Microsoft and has spent the past several years building regulatory data infrastructure for the pharmaceutical industry.
Kavita Pant is a pharmaceutical intelligence professional at Atlas Compliance with more than eight years of experience spanning regulatory analysis, competitive intelligence, market research, and drug profiling.
This content is sponsored by Atlas Compliance. The views and opinions expressed are those of the sponsor and do not necessarily reflect those of the Regulatory Affairs Professionals Society.