FDA, MHRA officials eye streamlined frameworks for regulating genAI
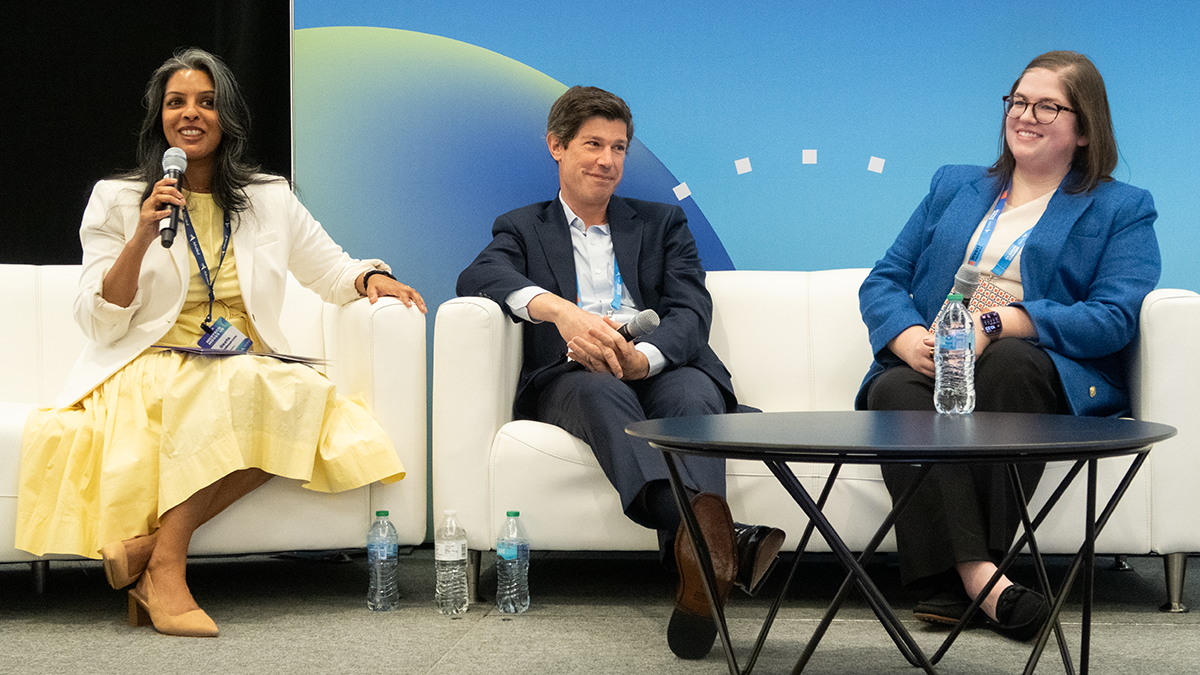
Geeta Pamidimukkala (left), Rick Abramson (center), and Jessica Paulsen (right) spoke at The MedTech Conference. (credit: Ferdous Al-Faruque)
Boston – Regulators from the US and UK discussed ongoing efforts to develop regulatory frameworks that are better suited to generative artificial intelligence (genAI) products at The MedTech Conference on Tuesday.
The officials said they want to help bring genAI-enabled products to market faster while also taking appropriate time to ensure they are not compromising their public health mission.
In recent months, the US Food and Drug Administration (FDA) and the UK Medicines and Healthcare products Regulatory Agency (MHRA) have published documents laying out their thinking on the regulation of various AI-enabled products. In August, FDA published a discussion paper laying out a competency-based framework to regulate genAI-enabled medical devices that would rely on non-clinical device benchmarking and clinical confirmation on how these devices perform. (RELATED: FDA seeks feedback on framework for regulating GenAI devices, Regulatory Focus 24 August 2026)
MHRA also adopted a list of 44 recommendations on the use of AI in medical devices developed by an independent group of doctors on the National Commission into the Regulation of AI in Healthcare. The recommendations are intended to create a proportionate regulatory framework and to emphasize a lifecycle approach for such products. (RELATED: MHRA announces strategy to streamline regulation in the AI era, Regulatory Focus 14 October 2026)
Rick Abramson, director of FDA's Digital Health Center of Excellence (DHCoE), stressed that the proposed competency-based framework for regulating genAI is not just an academic exercise, but rather the first step in developing draft policy on the topic. Abramson added that FDA has received hundreds of comments on the discussion paper and emphasized that AI is a priority at the highest levels of the Trump Administration.
"This is very, very important policy," he said. "In the course of months, expect draft policy guidance to be issued, and then there'll be another opportunity for stakeholder comment that we look forward to."
Panel moderator Geeta Pamidimukkala, vice president for technology and regulatory affairs at AdvaMed, noted that the Center for Devices and Radiological Health (CDRH) recently published its 2027 guidance priorities list, which includes two guidance documents specifically addressing genAI. She asked whether regulators were concerned about the speed at which guidances are developed and disseminated compared with how quickly genAI technology is evolving, and whether FDA is considering other ways to disseminate policy information outside the formal guidance process.
Abramson noted that the genAI discussion paper itself went through the administration's clearance process at a potentially record speed but also emphasized that regulators need to be thoughtful when developing policy. He acknowledged that stakeholders are concerned that while the technology evolves on a timeframe of weeks to months and guidance may take years, he is confident FDA will be able to adapt to the regulatory needs reasonably quickly.
"We want to move fast, but we don't want to move too fast," said Abramson. "This is very, very, very, very complex and very important policy, we're going to be setting precedent that lasts for years, and so we want to do it right.
"We could have moved directly to draft guidance, but we intentionally, very deliberately said, 'No, we were going to issue a discussion paper first,'" he added. "We want to engage the stakeholder community; we want to get input from various corners of the ecosystem to inform even our initial policy development efforts."
The panelists also discussed the use of predetermined change control plans (PCCPs) in the FDA discussion paper and the MHRA AI framework to enhance regulatory efficiency, particularly for devices that use genAI and for complex AI-enabled products. Pamidimukkala noted that MHRA has indicated it plans to issue a guidance on PCCP at the end of the year and asked Jessica Paulsen, deputy Director for AI and software at MHRA and former FDA acting director at DHCoE, how her experience at FDA may inform her work in helping MHRA develop the guidance.
Paulsen said that there are several documents that MHRA can build on, including FDA guidance on PCCPs; machine learning guiding principles developed by MHRA, FDA and Health Canada; as well as a PCCP technical document published by the International Medical Device Regulators Forum (IMDRF). She noted that the inclusion of PCCPs in the strategy document is an acknowledgment from the National Commission into the Regulation of AI in Healthcare that PCCPs can reduce submission burdens and increase patient access to products. She also said the commission has encouraged MHRA to consider the boundaries of genAI's performance capabilities as it considers how to regulate such products through PCCP.
"Another way to think about it is kind of like an envelope," said Paulsen. "How do you make sure that you can define the boundaries of that envelope so the device stays within it, remains safe, remains effective, and everyone has that assurance, but the device can iterate in a way that is efficient, and there's a clear path for it to support its safety.
"That's how we're thinking about it as we're drafting this guidance," she added. "It'll go out in draft, so we do want public comment on that."
Another area of the forthcoming PCCP guidance, according to Paulsen, is how MHRA can provide a clear path to genAI manufacturers to expand the use of their products. She noted it means taking into consideration the original indications for use while maintaining boundaries that limit how much the product can change.
Abramson said that even though FDA received explicit authority to allow PCCPs under the Food and Drug Omnibus Reform Act of 2022 (FDORA), the concept is still relatively new. He said the agency has received feedback requesting more clarification on how it will regulate products with PCCP and assured attendees it is continuously working to provide that feedback, including a document published the same day with examples of how PCCPs can be used.
Pamidimukkala also noted that both FDA and MHRA addressed the balance between pre- and postmarket oversight and the use of real-world data (RWD) in their documents. She asked how the regulators envisioned RWD contributing to their respective postmarket surveillance and how that could also inform their premarket evidentiary expectations.
Paulsen said that she is concerned that regulators may be over-relying on single-point-in-time assessments that may not accurately reflect how AI products perform in the real world. With that in mind, she noted that MHRA is thinking more holistically about what good postmarket monitoring looks like and how meaningful real-world evidence (RWE) and RWD can complement their total product lifecycle approach to reduce some premarket evidentiary requirements if they can get assurances that the products continue to function as intended.
Abramson emphasized that, in his personal view, he would like to see a robust data collection monitoring infrastructure in the postmarket setting that would allow for substantial balancing of evidentiary burdens between the premarket and postmarket setting. He noted that the discussion paper addresses this by asking whether a stronger, more robust postmarket data infrastructure could reduce premarket burdens.
Abramson also acknowledged that industry is likely to be skeptical of the government and argue that regulators would only increase postmarket burdens while keeping the premarket requirements.